

S. Korea overhauls regulatory framework to expand access to rare disease treatments
Nov 11, 2026, 10:20 am


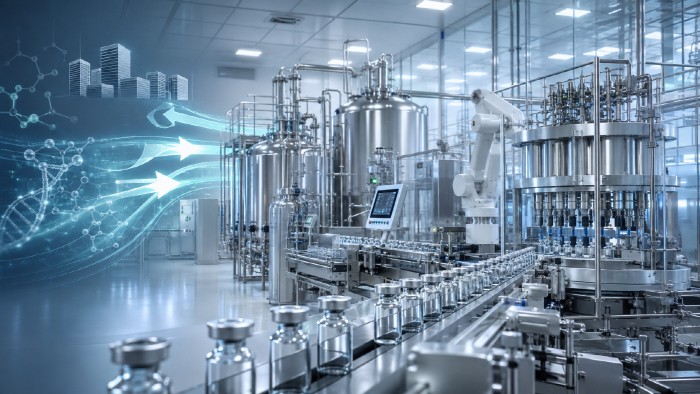 |
| Image for new drug development./AI-generated image |
South Korea’s Ministry of Food and Drug Safety is actively changing the drug supply system and new drug development standards to expand treatment opportunities for patients with rare and intractable diseases. Some treatments that patients had previously obtained directly from overseas will now be imported and supplied by public authorities, while guidelines have been prepared for designing clinical trials for diseases with too few patients to conduct studies easily. In addition, patient-reported quality of life may now be used as supplementary data in the marketing authorization review process.
According to the ministry on Nov. 10, the rapid introduction of orphan drugs was the first flagship task among the “50 food and drug safety initiatives” announced in November last year. The “60 initiatives” announced in July this year included clinical trial design for small patient groups and the use of patient experience in marketing authorization reviews.
A list of products converted to emergency introduction, released by the ministry in September, included 10 self-administered drugs such as tiopronin tablets. The ministry plans to convert at least 10 items a year in this way. A revised Pharmaceutical Affairs Act, set to take effect on Nov. 12, will allow the ministry to urgently import nationally essential medicines that are difficult to supply or commission pharmaceutical companies to produce them.
The designation criteria for orphan drugs were also revised to broaden the range of medicines available for use in Korea. In February, the ministry amended the relevant notification so that drugs used to treat or diagnose rare diseases under the Rare Disease Management Act can be designated on the basis of their intended use. This eased the previous burden of having to prepare comparative data showing that a new drug offered major improvements in safety or efficacy when an existing treatment was already available.
In early clinical trials for new anticancer drugs, participation had largely been limited to patients who had no remaining standard treatment options. In November last year, the ministry established review standards allowing patients with cancers that are difficult to cure to participate even before exhausting all standard treatments, based on consideration of the likely effectiveness of remaining treatments and the expected benefits and risks of the investigational drug.
For cell and gene therapies for rare diseases, recruiting participants is difficult from the clinical trial stage because patient numbers are so small. On Oct. 30, the ministry released guidelines outlining considerations for designing clinical trials involving a small number of patients. It also issued separate guidance on how to assess the quality of gene therapies for rare diseases and what should be confirmed before clinical trials begin.
Patient experience is also used in determining what changes to assess in clinical trials. On the same day, the ministry established the “guideline on marketing authorization review of pharmaceuticals based on patient experience data.” This makes it possible to use patients’ reported health status, quality of life, and treatment preferences from the early stages of development, and to use them at the approval stage as supplementary data in weighing a drug’s benefits and risks.
Products combining pharmaceuticals and digital medical devices will be reviewed by a joint team of reviewers from each field under the integrated review process established on Oct. 30. On Dec. 31, a special act supporting contract development and manufacturing organization (CDMO) companies in the biopharmaceutical sector in responding to export regulations will also take effect.
A ministry official said, “Based on thorough verification of safety and efficacy, the Ministry of Food and Drug Safety will do its utmost to support prompt approval reviews so that medicines essential for patient treatment can be supplied in a timely manner.”
Lee Se-mi
Most Read
-
1
Ukraine halts strikes on Russian oil refineries un..

-
2
China's fifth plenum to conclude amid expectations..

-
3
Southeast Asian auto parts makers hit hard by Chin..

-
4
U.S. links Myanmar peace efforts to rare earth acc..

-
5
Hegseth rejects calls for impeachment over Iran wa..

-
6
Democrats flip U.S. Congress in historic midterm s..

-
7
USFK commander cites interoperability gaps, urges..

PEOPLE


-
Address : In-Young Bldg., 34, Uisadang-daero 1-gil, Yeongdeungpo-gu, Seoul
Copyright by ASIATODAY CO., LTD. All rights resereved